AI & Materials
ISSN: 3006-7588 (Print)
ISSN: 3006-7596 (Online)
CODEN: AMABGR
Machine learning based prediction of luminescence properties of ion-doped Cs2ZrCl6 crystalline powders
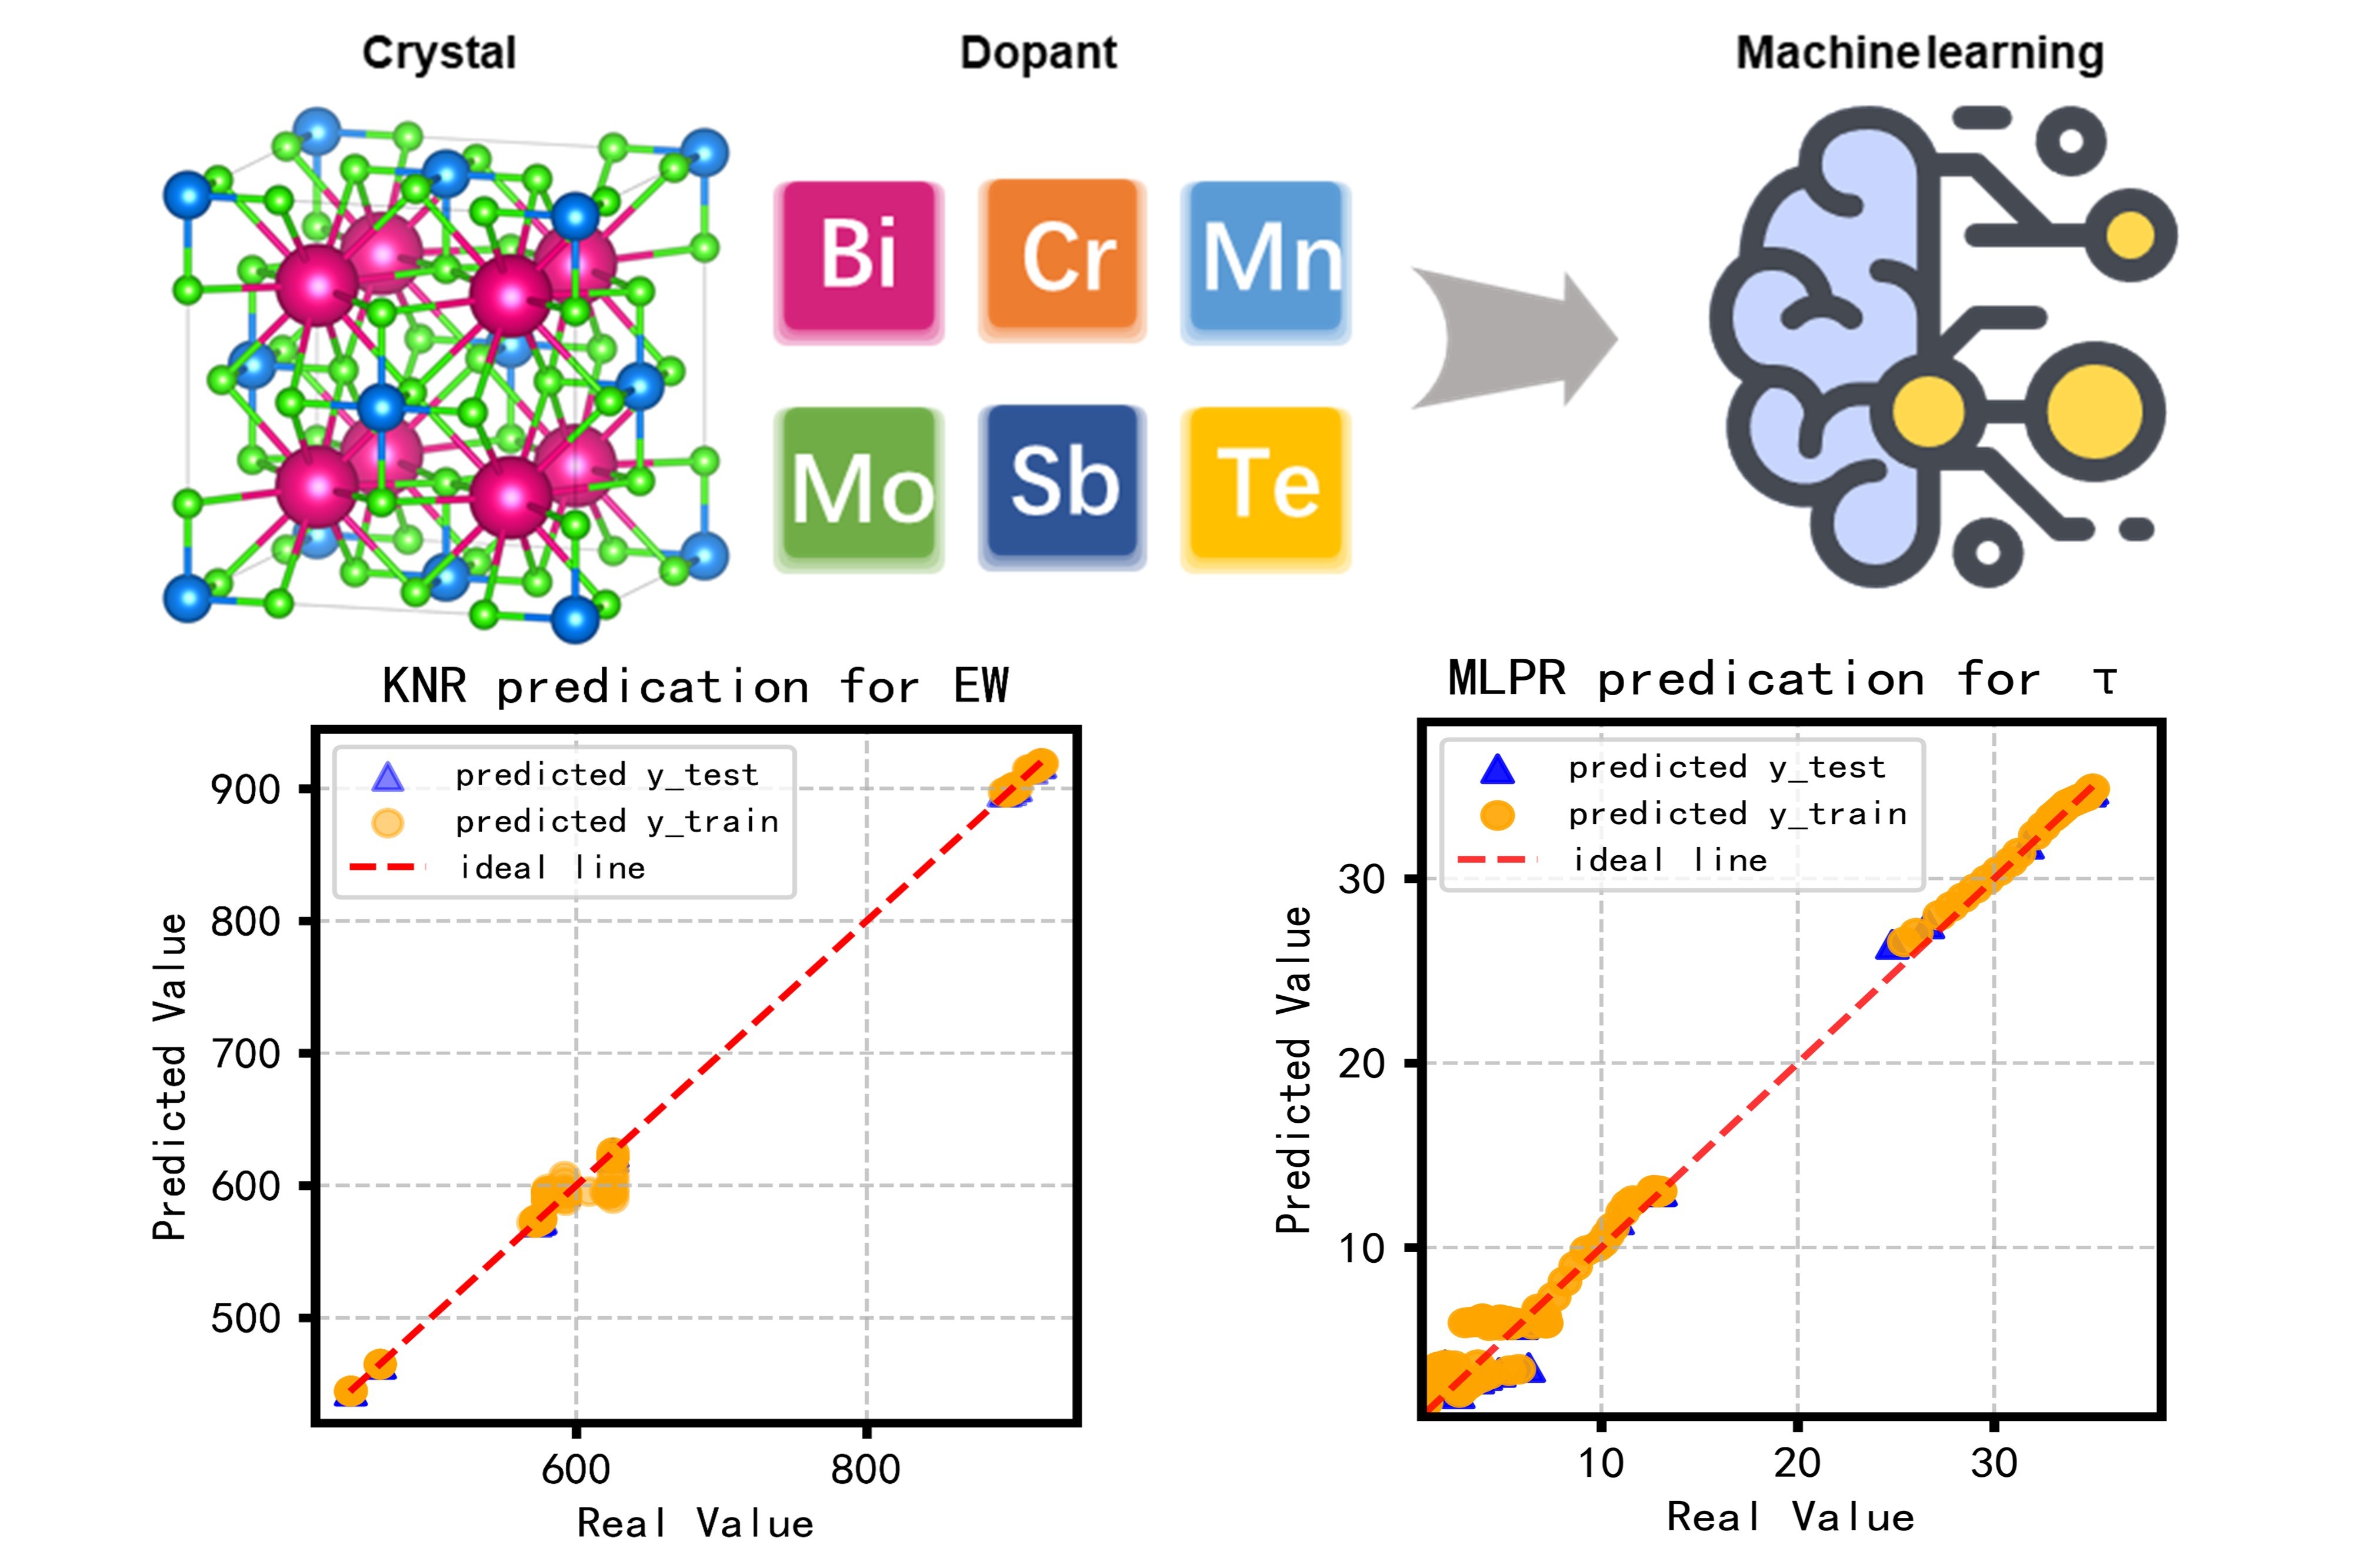
Predicting the luminescence behavior of Cs2ZrCl6 crystalline powders with speed and accuracy is a key prerequisite for designing new scintillators aimed at X-ray imaging. Conventional first-principles calculations based on density functional theory can reproduce crystal luminescence, yet they demand substantial computing power and deep materials-domain knowledge, which together translate into high cost, long development cycles, and limited throughput. To sidestep these bottlenecks, our work adopts a machine learning strategy to predict the luminescence properties of Cs2ZrCl6. A training dataset was assembled from the literature and expanded through linear interpolation. The hyperparameters of six regression algorithms, including ridge regression (RR), K-nearest neighbor regression (KNR), support vector regression (SVR), random forest regression (RFR), XGBoost regression (XGBR), and multilayer perceptron regression (MLPR), were tuned by coupling the covariance matrix adaptation evolution strategy (CMA-ES) with 10-fold cross-validation. Once trained, the models captured a reliable composition-structure-property link. By comprehensively evaluating three metrics of the coefficient of determination (R²), mean absolute error (MAE), and mean squared error (MSE), two optimized luminescence property prediction models were identified. The KNR model was selected for emission wavelength prediction, achieving a high R2 of 0.9999 and a low MSE of 1.4434. Meanwhile, the MLPR model was chosen for decay time prediction, with a high R2 of 0.9893 and a low MSE of 0.9234. Additionally, the Shapley Additive exPlanations (SHAP) analysis further quantified feature contributions to target properties, offering physically reasonable interpretations. This work demonstrates the promising capacity of machine learning for fast, reliable prediction of crystal luminescence performance, providing practical support for accelerating materials-oriented research and development.
Highly vertically oriented 2D/3D tin perovskite films for efficient and stable solar cells
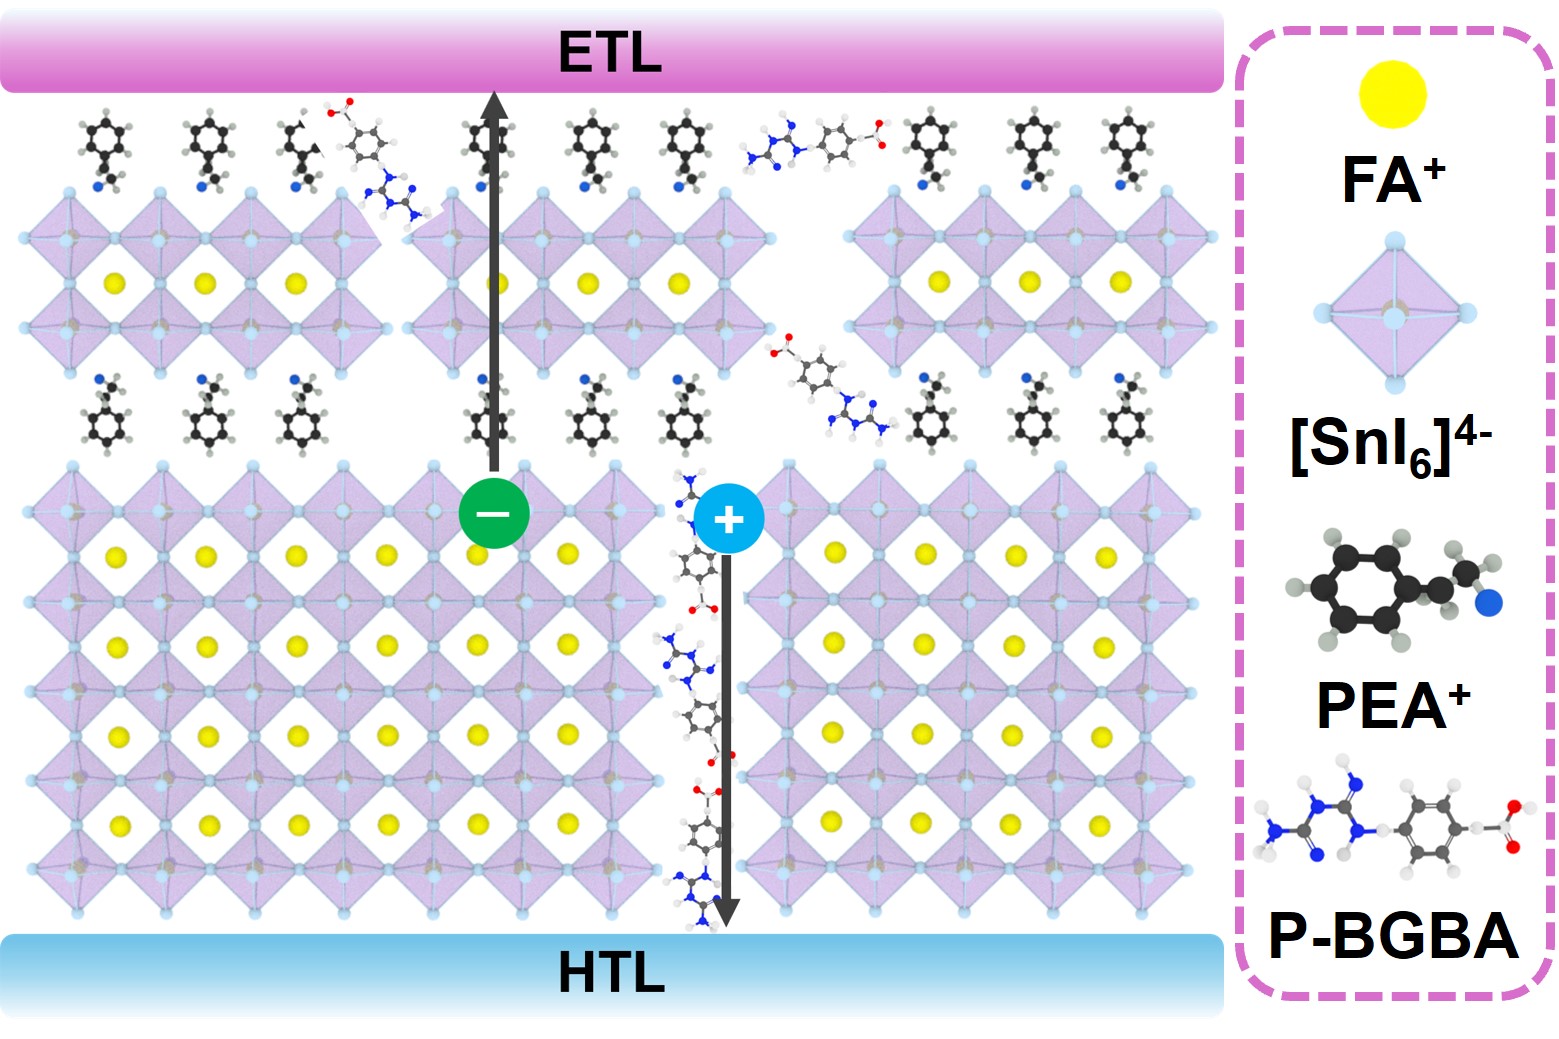
Two-dimensional/three-dimensional (2D/3D) tin perovskite solar cells (PSCs) have considerable potential for lead-free photovoltaics because of their low toxicity and favorable optoelectronic properties. However, their efficiency and stability remain limited by the difficulty of controlling the crystal orientation of 2D/3D perovskites. Here, a long-chain Lewis base molecule containing multiple functional groups, p-biguanidinobenzoic acid hydrochloride (p-BGBA), is introduced to regulate the crystallization kinetics of 2D/3D tin perovskites. The results show that p-BGBA effectively modulates the crystallization process through multiple interactions with the perovskite, thereby promoting the formation of high-quality films with a highly vertical crystal orientation. As a result, the resulting tin PSCs achieve a champion power conversion efficiency (PCE) of 14.96% and retain 91% of their initial performance after 2500 h of storage under a nitrogen atmosphere.
Machine learning-based ion sieve forming strategy: impact on salt lake lithium recovery by fixed beds
Lithium recovery from salt-lake brines is of great significance for addressing the growing demand for lithium resources. Fixed-bed adsorption is a key process in lithium recovery from salt lakes, and the shaping strategy of the adsorbent plays a decisive role in determining fixed-bed performance. This study is the first to apply machine learning to analyze the shaping strategy of fixed-bed adsorbents for lithium recovery from salt lakes. A multi-model analytical framework integrating back propagation (BP), radial basis function (RBF), probabilistic neural network (PNN), generalized regression neural network (GRNN), random forest (RF), and genetic algorithm (GA) models was established to evaluate the effects of material properties, process parameters, and shaping strategies on fixed-bed adsorption performance. The results showed that the liquid-film mass transfer efficiency has a greater influence on fixed-bed performance than the adsorbent material itself. The optimal structural parameters were identified using the radial basis function–genetic algorithm (RBF-GA) model and further validated experimentally by response surface methodology (RSM). The optimized conditions were determined to be a packing diameter of 12.5 mm and a packing density of 0.30. This work provides new insight into the structural design of fixed-bed adsorbents and offers a useful reference for the development and optimization of lithium recovery processes from salt-lake brines.

Research progress on SiCf/SiC composites and their cladding components
Owing to their outstanding high-temperature resistance, oxidation resistance, radiation tolerance, and corrosion resistance, silicon carbide fiber-reinforced silicon carbide (SiCf/SiC) composites have shown extensive potential in advanced applications, including aerospace and nuclear industries. SiCf/SiC composites are considered among the most promising accident-tolerant fuel cladding materials, especially in the context of fourth-generation fission reactor development, owing to their stability under extreme conditions. However, the complex processing and structural characteristics of the material leave room for further research, especially with the emergence of new technologies like artificial intelligence (AI). Therefore, this work will provide a review of various processes, including chemical vapor infiltration (CVI), polymer infiltration and pyrolysis (PIP), nano impregnation and transient eutectic method (NITE), and reactive melt infiltration (RMI), focusing on improving material density, mechanical properties, and irradiation stability. Additionally, an in-depth review of the mechanical properties and microstructural changes of SiCf/SiC composites and their cladding components under extreme conditions, such as high temperatures, irradiation, and corrosion, is provided, as these factors directly affect their long-term stability in nuclear reactors. Notably, numerical simulation technology has become a crucial tool for predicting the service performance of materials. Integrating advanced technologies like AI is expected to further promote the application of SiCf/SiC composites in future high-temperature structural materials. In summary, significant progress has been made in the study of SiCf/SiC composites as next-generation nuclear fuel cladding materials. However, further research is needed in areas such as fabrication process optimization, interface modification, service behavior evaluation, and integration with AI to meet the higher performance demands of future nuclear energy systems.

Applications of artificial intelligence in materials research for fuel cells
The increasing global energy demand and the growing environmental problems have intensified the pursuit of clean and sustainable energy solutions. Hydrogen, with its high energy density and clean by-products, is a promising candidate as an energy source. Fuel cells play a key role in harnessing hydrogen energy, but this technology faces challenges such as the trade-off between material stability and ion conductivity, which limits its widespread application. To address these challenges, designing material properties and adjusting system parameters are highly desirable. However, the traditional trial-and-error approach is no longer feasible when dealing with the vast array of possibilities. Fortunately, the advancement of artificial intelligence (AI) offers a new approach which can dramatically speed up the material design and parameter control. This article reviews the application of AI in fuel cells, especially its ability to accelerate material development. The review begins by outlining the mechanisms and classifications of fuel cells, as well as the property requirements for each part of the fuel cells. Subsequently, the article introduces the basic concepts of AI and its application in materials science, including the workflows of data aggregation, feature construction, model training, and experimental validation. Importantly, the applications of AI in predicting fuel cell material performance are highly emphasized and discussed. In addition, the challenges encountered in AI applications are introduced, including sparse datasets, complex feature engineering, the limitations of general models, and the weak interpretability of AI models, along with their respective development blueprints.
Materials discovery through reinforcement learning: a comprehensive review
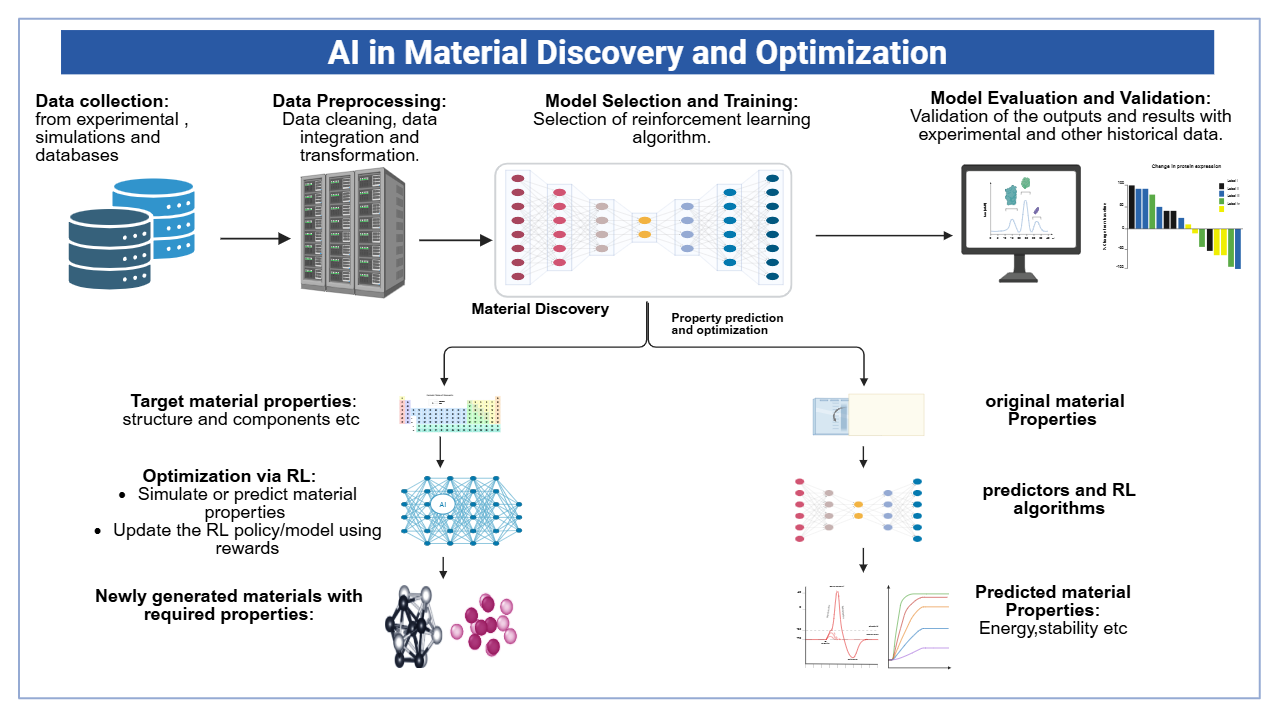
Reinforcement learning (RL) is emerging as a powerful tool in materials science, delivering a paradigm shift in how we find and optimize high-dimensional chemical and structural spaces. Unlike traditional methods, RL agents are able to learn to explore complex energy landscapes in an adaptive manner, instantaneously making decisions that guide the discovery of novel materials with certain properties. However, the application of RL to materials discovery faces unique challenges, including data scarcity, computationally expensive, and the challenge of designing reward functions that can balance multiple material objectives optimally. In this review, the current challenges and difficulties in applying RLtechniques in materials science and recent advances combining RL with machine learning, generative models, and domain knowledge are emphasized. We also outline promising future directions, such as transfer learning, hybrid models, and the creation of collaborative, open-access data infrastructures. By addressing these challenges, RL has the potential to transform the discovery and design of functional materials for catalysis, energy storage, and sustainability applications.